iTranslated by AI
[Single-cell Analysis] Seurat: Cell Subsetting Examples using subset()



After performing a rough analysis of cell data obtained from single-cell analysis with Seurat, it is common to extract specific cell populations for more detailed analysis. From a Seurat object, cells can be extracted using index operations or the subset() function. I have summarized some small tips here.
Table of Contents
- Extracting by active.ident information
- Extracting by meta.data columns
- Extracting by expression levels
- Extracting by cell barcodes
- Extraction by UMAP values
- Cell extraction by mouse operation
Demo data
Preparing the data
Using the PBMC 2,700 cell data available from the Seurat tutorial page.
Processing proceeds in the same way as the tutorial. For demonstration purposes, cluster number information and cell name information are stored in the meta.data slot.
library(Seurat)
library(dplyr)
pbmc.data <- Read10X(data.dir = "pbmc3k_filtered_gene_bc_matrices/filtered_gene_bc_matrices/hg19")
pbmc <- CreateSeuratObject(counts = pbmc.data, project = "pbmc3k", min.cells = 3, min.features = 200)
pbmc[["percent.mt"]] <- PercentageFeatureSet(pbmc, pattern = "^MT-")
pbmc <- subset(pbmc, subset = nFeature_RNA > 200 & nFeature_RNA < 2500 & percent.mt < 5)

pbmc <- pbmc %>%
  NormalizeData() %>%
  FindVariableFeatures() %>%
  ScaleData() %>%
  RunPCA() %>%
  RunUMAP(dims = 1:10) %>%
  FindNeighbors(dims = 1:10) %>%
  FindClusters(resolution = 0.5)
pbmc$Cluster <- pbmc@active.ident
DimPlot(pbmc, reduction = "umap", label = TRUE, pt.size = 0.5) + NoLegend()

new.cluster.ids <- c("Naive CD4 T", "CD14+ Mono", "Memory CD4 T", "B", "CD8 T", "FCGR3A+ Mono",
    "NK", "DC", "Platelet")
names(new.cluster.ids) <- levels(pbmc)
pbmc <- RenameIdents(pbmc, new.cluster.ids)
pbmc$Cell_type <- pbmc@active.ident
DimPlot(pbmc, reduction = "umap", label = TRUE, pt.size = 0.5, repel = T) + NoLegend()

Extracting by active.ident information
active.ident is the information stored in seurat_object@active.ident, which temporarily holds the cluster information currently being used.
Various plotting functions and features such as FindMarkers() and AverageExpression() also use the active.ident information unless specified otherwise.
active.ident can be changed simply by specifying the column name of the meta.data slot as follows.
Idents(pbmc) <- "Cluster"
pbmc <- SetIdent(pbmc, value = "Cluster")
pbmc@active.ident <- pbmc@meta.data$Cluster
To extract cells based on active.ident information, use the idents= argument.
The return value of subset() is a Seurat object.
When active.ident is cluster numbers
pbmc2 <- subset(pbmc, idents = 0)
DimPlot(pbmc2)

For cluster numbers, you can use them with or without quotation marks.
pbmc2 <- subset(pbmc, idents = "1")
DimPlot(pbmc2)

You can extract multiple clusters at once using a vector of cluster numbers.
pbmc2 <- subset(pbmc, idents = c(2,3,4))
DimPlot(pbmc2)

When active.ident is text such as cell names
Idents(pbmc) <- "Cell_type"
In the case of strings, specify them with quotation marks. You can also extract multiple items at once here as well.
pbmc2 <- subset(pbmc, idents = c("Naive CD4 T","B"))
DimPlot(pbmc2)

Extracting everything except the specified items (invert)
By setting the invert= argument to TRUE, you can extract cells other than those specified.
pbmc2 <- subset(pbmc, idents = c("Naive CD4 T","B"), invert = TRUE)
DimPlot(pbmc2)

Extracting by meta.data columns
To extract cells using meta.data columns instead of active.ident, use the subset= argument within the subset() function.
Write it like subset(seurat_object, subset = column_name == "desired_factor").
The column name is specified without quotation marks.
pbmc2 <- subset(pbmc, subset = Cluster == "1")
DimPlot(pbmc2)

Extracting by expression levels
It is also possible to extract cells based on specific gene expression. In this case, too, do not use quotation marks for gene names.
pbmc2 <- subset(pbmc, subset = CD4 > 1)
DimPlot(pbmc2)

Multiple conditions can also be combined.
pbmc2 <- subset(pbmc, subset = CD3E>1 & CD4>1)
DimPlot(pbmc2)
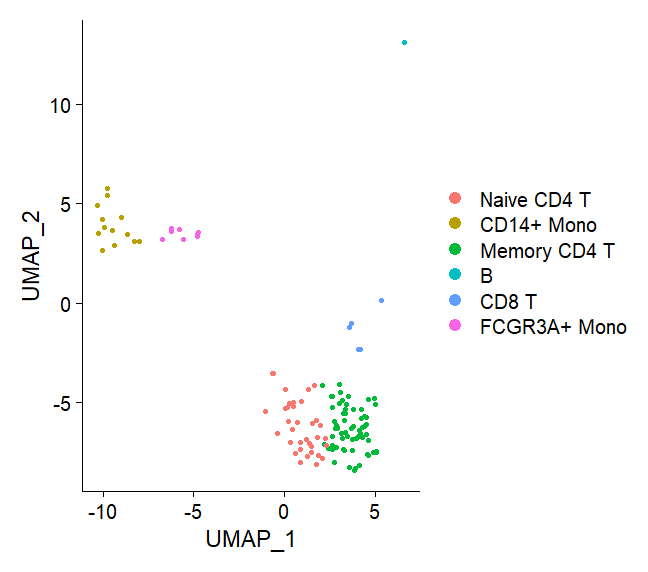
In Seurat, there are three types of expression data in the assay slot: (1) counts, (2) data, and (3) scale.data.
The slot can be specified with slot=. By default, it seems to refer to the data slot.
pbmc2 <- subset(pbmc, subset = CD3E>1 & CD4>1, slot = "counts")
DimPlot(pbmc2)

If there are multiple assays, it is safer to change the DefaultAssay before proceeding.
Extracting by cell barcodes
If you have prepared a vector of cell barcodes under specific conditions, it is also possible to extract cells using the cells= argument.
cells <- WhichCells(object = pbmc, idents = "Naive CD4 T")

pbmc2 <- subset(pbmc, cells = cells)
DimPlot(pbmc2)

Extraction by UMAP values
When performing dimensional reduction, small cell populations that look like outliers may sometimes be generated.
Even if they can be distinguished by a different cluster number from other populations through clustering, it doesn't always go that conveniently.
If they are sufficiently separated from other populations on UMAP, you can also extract cells using the values of the 1st and 2nd dimensions of UMAP.
For example, in this UMAP, the B cell population is located separately in the upper right, and it seems possible to separate them using only the values of the 2nd dimension of UMAP.
DimPlot(pbmc)

pbmc2 <- subset(pbmc, subset = UMAP_2 < 10)
DimPlot(pbmc2)

Extract the lower right part
pbmc2 <- subset(pbmc, subset = UMAP_1 > -1 & UMAP_2 < 3)
DimPlot(pbmc2)

Cell extraction by mouse operation
Passing the output of DimPlot() to CellSelector() launches a viewer that allows you to enclose cell populations using mouse operations, powered by the shiny package.
plot <- DimPlot(pbmc)
select.cells <- CellSelector(plot = plot)

Drag on the screen to enclose the desired cell population. Press Done to finish.

The return value of CellSelector is a vector of cell barcodes.

It can be used by passing it to the cells= argument.
pbmc2 <- subset(pbmc, cells = select.cells)
DimPlot(pbmc2)

Discussion